Home / Small molecules / Drug product / Oral drug products / Formulation development
Formulation development
The formulation that goes into Phase III is the one that will be filed.
Early phase formulation development
Our oral solid formulation development teams work on drug candidates from preclinical through clinical to commercial manufacturing, and have tackled poor bioavailability, excipient incompatibility, vehicle stability and release profile.
Formulation development typically takes 1 to 6 weeks. From GMP API received to Phase I clinical trial material released is 4 to 6 weeks.
High-potency oral formulations are developed and manufactured at an occupational exposure limit of 10 ng/m³.
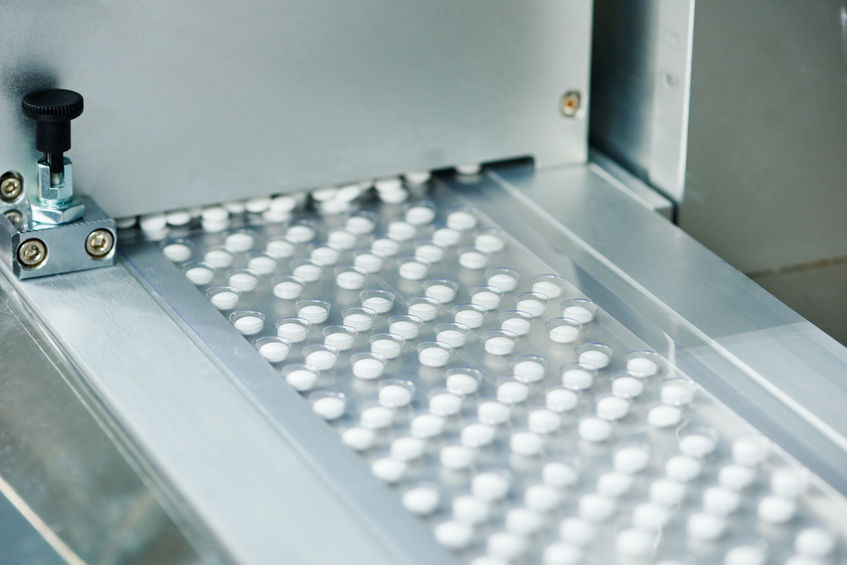
Flexible formulation and process options
- API in bottle or capsule
- Powder in capsule
- Tablets
- Lipid formulation
- Roller compaction
- Direct compression
- Wet granulation
Bioavailability enhancement
A comprehensive enabling technology platform covers spray dried dispersion, hot melt extrusion, nano suspension and lipid formulation, with lipid nanoparticle available on the parenteral platform.
The candidates are screened in parallel rather than selected on paper, because a technology that will fail for a molecule usually only reveals that months into development.
Fast to Clinic Supply
F2CS is a service package for programs approaching IND submission: formulation development and clinical trial material supply both within 8 weeks.
Integrated drug substance, drug product, analytical and CMC writing teams work in parallel. Drug product and analytical teams sit within a two-hour drive of the drug substance teams, which is what makes material transfer and scheduling quick.
The package draws on experience across 1,000+ preclinical to Phase III molecules a year in a range of dosage forms.
Late phase formulation development
The Phase I formulation is usually the simplest one that can deliver the dose, chosen on limited API and limited stability data. Between Phase I and Phase III it is developed into the formulation that will be filed: the dosage form and strengths the clinical dose now supports, excipient grades and levels set against compatibility and process data, a manufacturing process chosen on the API’s flow, compressibility and moisture sensitivity, and a dissolution method that discriminates between variants.
Most of that work is driven by manufacturability and stability. Changes to tablet size, dosing frequency or a modified-release profile are settled as early as possible, because each can change exposure and needs a clinical bridge. The formulation is locked before pivotal dosing wherever the program allows.
Late phase teams take candidates through pivotal clinical supply, process optimization by design of experiments, process validation and NDA registration.
Fast for Commercial Launch
F4CL is the late-phase package: technology transfer to release of PPQ completed in 6 months.
Formulation and analytical teams across North America, Europe and Asia support global filings, which have different requirements and timelines by agency. GMP facilities in Shanghai, Wuxi City and Couvet have been inspected by the US FDA, EMA, China NMPA, Japan PMDA, SwissMedic and other agencies, and are equipped for spray dried dispersion, hot melt extrusion, dry, wet and fluid bed granulation, continuous manufacturing and high potency drug product.
Managing change between IND and validation
Between the IND formulation and the validated one, most programs make changes. Each change carries a stability obligation and a comparability question, so the aim is to make as few as possible while still arriving at a manufacturable process.
Deciding which change justifies a bridging study, and which does not, is where prior late-phase experience is applied.